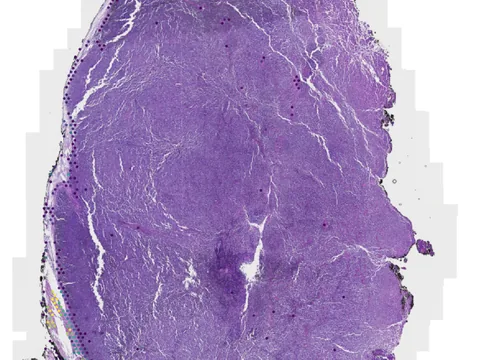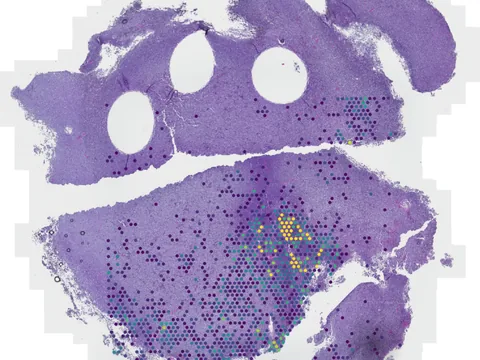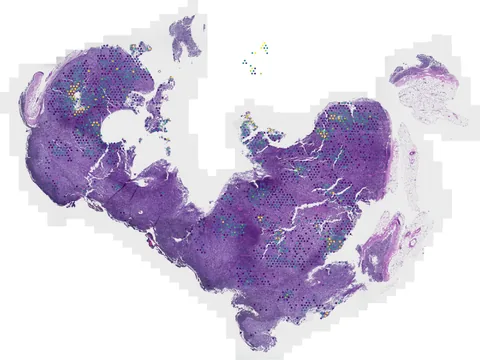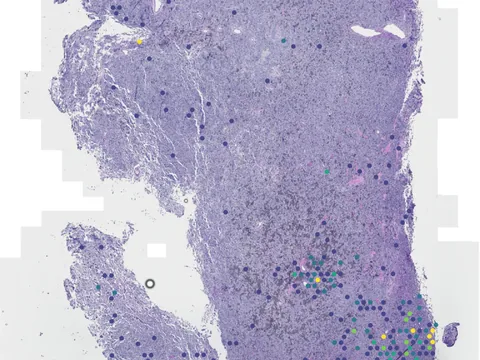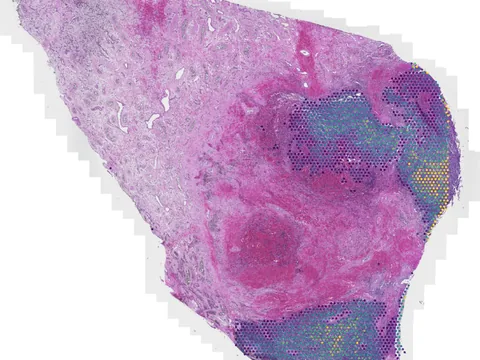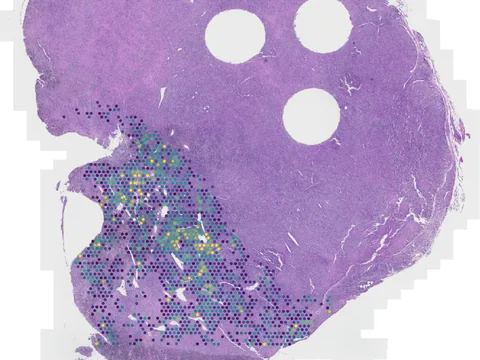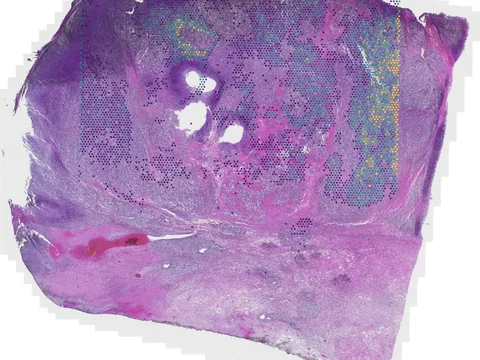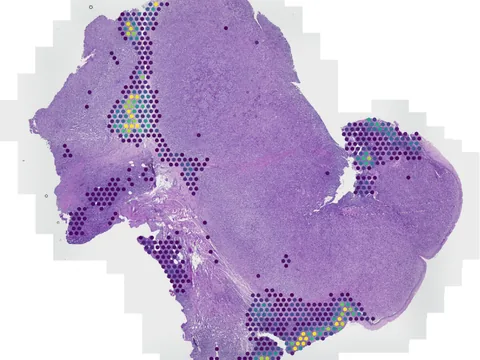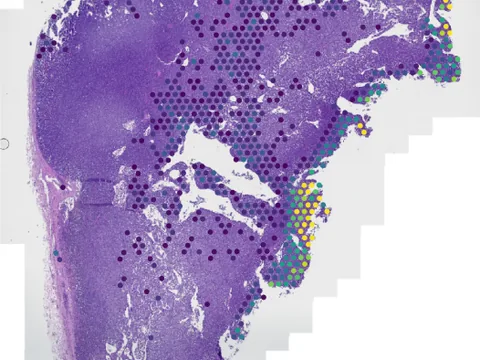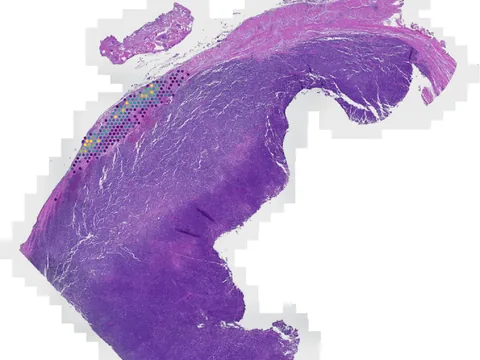
Spatial transcriptomics of HIV-associated non-Hodgkin lymphomas by EBV status and cell of origin
Non-Hodgkin Lymphoma (NHL) is the main cancer-related mortality for people living with HIV (PLWH), who have elevated risk for lymphoma relative to the general population. NHL genetic and molecular classifications have been intensely studied and correlated with clinical outcomes, but critical unanswered questions relevant to malignancy persist. For example, tumors positive for Epstein-Barr virus (EBV) account for 30-50% of HIV-associated NHLs, are particularly aggressive, and have poorer clinical outcomes – yet insights on the nature of EBV in NHL pathogenesis or potential therapeutic vulnerabilities have been limited. Here, we examined HIV-associated NHLs stratified by cell of origin (COO), molecular subtype, and EBV status using genome-wide spatial transcriptomic analyses.
import lamindb as ln
import spatialdata as sd
coll = ln.Collection.get(key="geo-gse274051")
coll.ordered_artifacts.df() # the 10 members, ordered
# download every member .zarr to the local cache:
paths = [a.cache() for a in coll.ordered_artifacts.all()]
# …or load each straight into memory as a SpatialData object:
sdatas = {a.key: sd.read_zarr(a.cache()) for a in coll.ordered_artifacts.all()} Cite this collection
No linked publication found — please cite the data source.